
Beyond the scientific and epidemiological reasons to doubt the lab escape theory of pandemic origin, let’s not forget that no US or allied intelligence has emerged with even a whiff of evidence that a lab was the origin by accident or intention.
https://t.co/vaZ5fY9UhD
https://t.co/TpS6tQppsS
https://t.co/TpS6tQppsS
https://t.co/twqUBqUNq9
https://t.co/LJ4tG6X47q
https://t.co/BZpvjfxsxK
https://t.co/LwFbHv9n1M
https://t.co/fsSxnJfJQQ
https://t.co/Qeu8jBblhV
More from Science




Hard agree. And if this is useful, let me share something that often gets omitted (not by @kakape).
Variants always emerge, & are not good or bad, but expected. The challenge is figuring out which variants are bad, and that can't be done with sequence alone.
You can't just look at a sequence and say, "Aha! A mutation in spike. This must be more transmissible or can evade antibody neutralization." Sure, we can use computational models to try and predict the functional consequence of a given mutation, but models are often wrong.
The virus acquires mutations randomly every time it replicates. Many mutations don't change the virus at all. Others may change it in a way that have no consequences for human transmission or disease. But you can't tell just looking at sequence alone.
In order to determine the functional impact of a mutation, you need to actually do experiments. You can look at some effects in cell culture, but to address questions relating to transmission or disease, you have to use animal models.
The reason people were concerned initially about B.1.1.7 is because of epidemiological evidence showing that it rapidly became dominant in one area. More rapidly that could be explained unless it had some kind of advantage that allowed it to outcompete other circulating variants.
Variants always emerge, & are not good or bad, but expected. The challenge is figuring out which variants are bad, and that can't be done with sequence alone.
Feels like the next thing we're going to need is a ranking system for how concerning "variants of concern\u201d actually are.
— Kai Kupferschmidt (@kakape) January 15, 2021
A lot of constellations of mutations are concerning, but people are lumping together variants with vastly different levels of evidence that we need to worry.
You can't just look at a sequence and say, "Aha! A mutation in spike. This must be more transmissible or can evade antibody neutralization." Sure, we can use computational models to try and predict the functional consequence of a given mutation, but models are often wrong.
The virus acquires mutations randomly every time it replicates. Many mutations don't change the virus at all. Others may change it in a way that have no consequences for human transmission or disease. But you can't tell just looking at sequence alone.
In order to determine the functional impact of a mutation, you need to actually do experiments. You can look at some effects in cell culture, but to address questions relating to transmission or disease, you have to use animal models.
The reason people were concerned initially about B.1.1.7 is because of epidemiological evidence showing that it rapidly became dominant in one area. More rapidly that could be explained unless it had some kind of advantage that allowed it to outcompete other circulating variants.

You May Also Like

https://t.co/6cRR2B3jBE
Viruses and other pathogens are often studied as stand-alone entities, despite that, in nature, they mostly live in multispecies associations called biofilms—both externally and within the host.
https://t.co/FBfXhUrH5d

Microorganisms in biofilms are enclosed by an extracellular matrix that confers protection and improves survival. Previous studies have shown that viruses can secondarily colonize preexisting biofilms, and viral biofilms have also been described.

...we raise the perspective that CoVs can persistently infect bats due to their association with biofilm structures. This phenomenon potentially provides an optimal environment for nonpathogenic & well-adapted viruses to interact with the host, as well as for viral recombination.

Biofilms can also enhance virion viability in extracellular environments, such as on fomites and in aquatic sediments, allowing viral persistence and dissemination.

Viruses and other pathogens are often studied as stand-alone entities, despite that, in nature, they mostly live in multispecies associations called biofilms—both externally and within the host.
https://t.co/FBfXhUrH5d
Microorganisms in biofilms are enclosed by an extracellular matrix that confers protection and improves survival. Previous studies have shown that viruses can secondarily colonize preexisting biofilms, and viral biofilms have also been described.
...we raise the perspective that CoVs can persistently infect bats due to their association with biofilm structures. This phenomenon potentially provides an optimal environment for nonpathogenic & well-adapted viruses to interact with the host, as well as for viral recombination.
Biofilms can also enhance virion viability in extracellular environments, such as on fomites and in aquatic sediments, allowing viral persistence and dissemination.

Keep dwelling on this:
Further Examination of the Motif near PRRA Reveals Close Structural Similarity to the SEB Superantigen as well as Sequence Similarities to Neurotoxins and a Viral SAg.
The insertion PRRA together with 7 sequentially preceding residues & succeeding R685 (conserved in β-CoVs) form a motif, Y674QTQTNSPRRAR685, homologous to those of neurotoxins from Ophiophagus (cobra) and Bungarus genera, as well as neurotoxin-like regions from three RABV strains
(20) (Fig. 2D). We further noticed that the same segment bears close similarity to the HIV-1 glycoprotein gp120 SAg motif F164 to V174.
https://t.co/EwwJOSa8RK
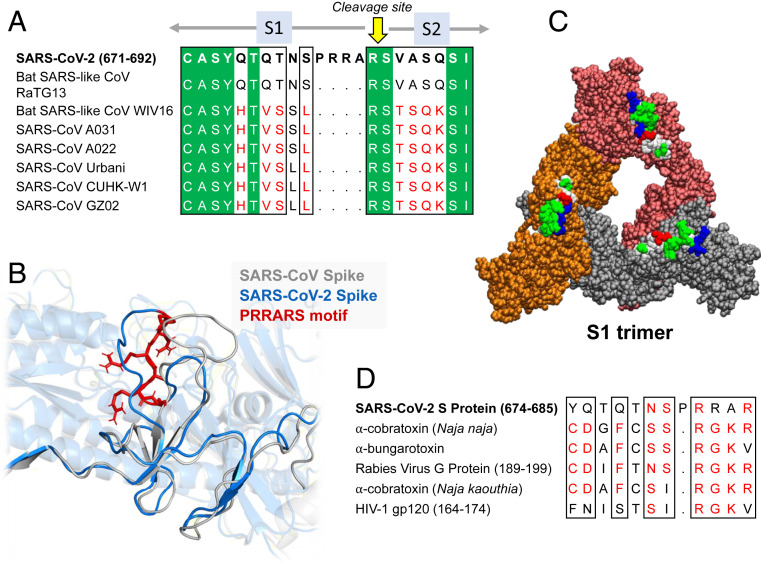
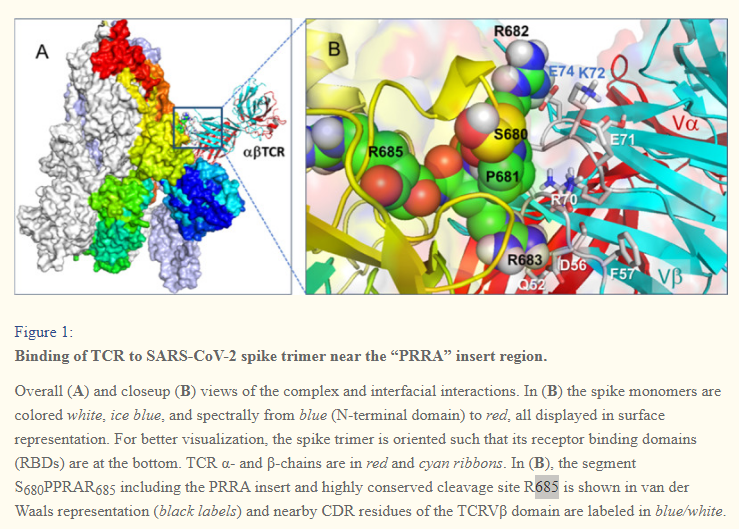
In (B), the segment S680PPRAR685 including the PRRA insert and highly conserved cleavage site *R685* is shown in van der Waals representation (black labels) and nearby CDR residues of the TCRVβ domain are labeled in blue/white
https://t.co/BsY8BAIzDa
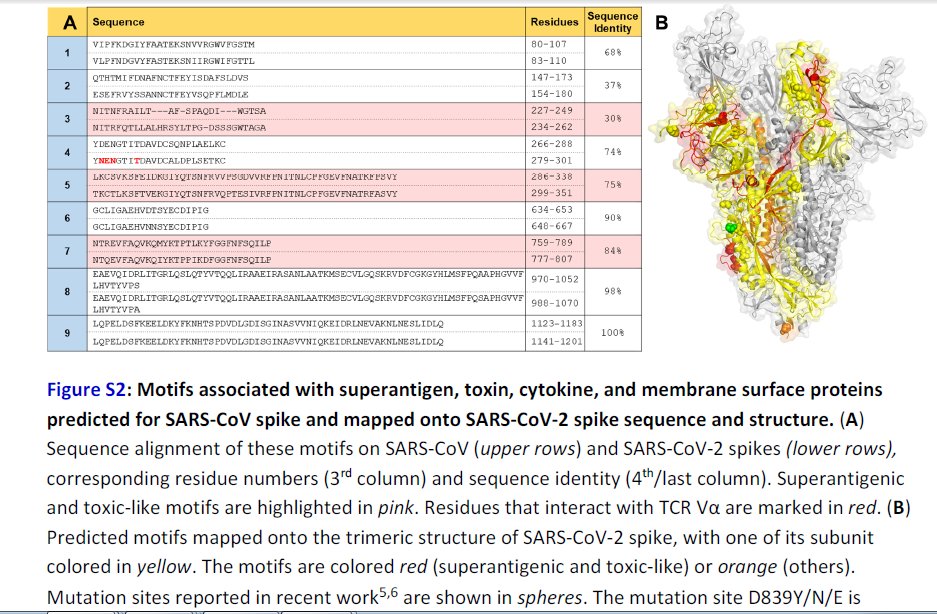
Sequence Identity %
https://t.co/BsY8BAIzDa
Y674 - QTQTNSPRRA - R685
Similar to neurotoxins from Ophiophagus (cobra) & Bungarus genera & neurotoxin-like regions from three RABV strains
T678 - NSPRRA- R685
Superantigenic core, consistently aligned against bacterial or viral SAgs
Further Examination of the Motif near PRRA Reveals Close Structural Similarity to the SEB Superantigen as well as Sequence Similarities to Neurotoxins and a Viral SAg.
The insertion PRRA together with 7 sequentially preceding residues & succeeding R685 (conserved in β-CoVs) form a motif, Y674QTQTNSPRRAR685, homologous to those of neurotoxins from Ophiophagus (cobra) and Bungarus genera, as well as neurotoxin-like regions from three RABV strains
(20) (Fig. 2D). We further noticed that the same segment bears close similarity to the HIV-1 glycoprotein gp120 SAg motif F164 to V174.
https://t.co/EwwJOSa8RK
In (B), the segment S680PPRAR685 including the PRRA insert and highly conserved cleavage site *R685* is shown in van der Waals representation (black labels) and nearby CDR residues of the TCRVβ domain are labeled in blue/white
https://t.co/BsY8BAIzDa
Sequence Identity %
https://t.co/BsY8BAIzDa
Y674 - QTQTNSPRRA - R685
Similar to neurotoxins from Ophiophagus (cobra) & Bungarus genera & neurotoxin-like regions from three RABV strains
T678 - NSPRRA- R685
Superantigenic core, consistently aligned against bacterial or viral SAgs


